MEDICINAL CHEMISTRY AT ITS BEST, Tracks information on drugs on worldwide basis by Dr Anthony Melvin Crasto, worlddrugtracker, helping millions with websites, 6 million hits on google, one lakh connections worldwide, email amcrasto@gmail.com, call +91 9323115463 India
antiviral, HEC 110114; Yiqibuvir, CHINA 2025, APPROVALS 2025, 82E4Q8WQV7
Encofosbuvir is a novel, oral small-molecule direct-acting antiviral (DAA) drug used to treat the hepatitis C virus (HCV). Approved by China's National Medical Products Administration (NMPA) in March 2025, it serves as a core component of a domestic, pan-genotypic treatment regimen.
🔬 Mechanism of Action
Encofosbuvir works by targeting the machinery the virus needs to replicate itself:
Target Enzyme: It functions as an HCV NS5B RNA-dependent RNA polymerase inhibitor.
Viral Suppression: By selectively binding to this polymerase, it blocks the synthesis of viral RNA, effectively halting the replication and spread of the hepatitis C virus in mammals
Clinical Indications and Usage
According to the official regulatory updates from the China NMPA, encofosbuvir is prescribed under specific clinical parameters:
Combination Therapy: It must be used in combination with netanasvir (specifically netanasvir phosphate capsules).
Target Genotypes: The regimen is highly effective across multiple viral strains, covering HCV genotypes 1, 2, 3, and 6.
Patient Profile: It is indicated for adult patients who are treatment-naïve (never treated before) or who have been previously treated with interferon. It is safe for use in patients with or without compensated liver cirrhosis.
The drug is classified as a Class 1 innovative drug, representing a milestone in self-developed, domestic intellectual property:
Developers: It was jointly developed and brought to market by Sunshine Lake Pharma (a subsidiary of HEC Pharm) and YiChang HEC ChangJiang Pharmaceutical Co., Ltd.
Dosage Form: It is distributed commercially as 0.3g tablets.
Therapeutic Context: This medication expands the developer's innovative hepatitis C pipeline, building upon their previously approved portfolio like emitasvir phosphate
OriginatorHEC Pharm; Sunshine Lake Pharma
DeveloperSunshine Lake Pharma
ClassAntivirals
Mechanism of ActionHepatitis C virus NS 5 protein inhibitors
RegisteredHepatitis C
27 Mar 2025Registered for Hepatitis C (Combination therapy, Treatment-naive) in China (PO)
08 Feb 2025Preregistered for Hepatitis C (Combination therapy, Treatment-experienced) in China (PO)
08 Feb 2025Registered for Hepatitis C (Combination therapy, Treatment-experienced) in China (PO)
Encofosbuvir is an antiviral drug used to treat hepatitis C virus (HCV). In China, encofosbuvir is approved for use in combination with netanasvir for the treatment of adult patients with chronic HCV genotypes 1, 2, 3, or 6, who are either treatment-naive or have been previously treated with interferon.[1]
Compound 3-6 (3.63 g, 4.2 mmol, 1 eq) was dissolved in acetone (12 mL), and water (9 mL), trifluoroacetic acid (3 mL), and glacial acetic acid (12 mL) were added sequentially at room temperature, followed by a reaction time of 2 hours. After the reaction was monitored by TLC until complete, 30 mL of dichloromethane was added to the reaction solution, stirred thoroughly, and allowed to stand for phase separation. The organic phase was washed sequentially with water (10 mL × 3), saturated sodium chloride (10 mL), dried over anhydrous sodium sulfate, and the solvent was removed under reduced pressure. The solution was purified by column chromatography using DCM:MeOH = 50:1 as the eluent, yielding 2.8 g of a white foamy solid.
Engasertib is an oral, once-daily AKT inhibitor developed by Vaderis Therapeutics, primarily investigated as a targeted therapy for Hereditary Hemorrhagic Telangiectasia (HHT). Clinical trials show it safely reduces the frequency and duration of bleeding episodes without an FDA-approved equivalent currently available
Core Information
Mechanism of Action: Engasertib is a highly selective inhibitor of AKT1 and AKT2. In HHT, mutations in the ALK1 pathway lead to abnormal blood vessel growth driven by an excess of the AKT protein. By inhibiting AKT, the drug promotes vascular stability and reduces vessel fragility.
Target Indication: Hereditary Hemorrhagic Telangiectasia (HHT) — a rare, severe genetic disorder causing vascular abnormalities and frequent, heavy bleeding, particularly nosebleeds (epistaxis)
Clinical Efficacy & Safety
Proof-of-Concept Trial: A 12-week, placebo-controlled study with 75 HHT patients evaluated daily doses of 30 mg and 40 mg.
The 40 mg cohort demonstrated a 41% reduction in mean bleeding duration and a 28% reduction in bleeding frequency, compared to 24% and 18% in the placebo group.
61% of patients in the 40 mg group rated their clinical condition as "Much Better".
Extended Efficacy: In long-term open-label extensions, benefits were sustained and amplified over 12 months, resulting in a 66% reduction in mean bleeding duration and a 55% reduction in bleeding frequency.
Side Effects: Generally well-tolerated. The most common side effects (reversible and manageable with supportive care) were mild-to-moderate rash and hyperglycemia
OriginatorAlmac Discovery
DeveloperVaderis Therapeutics
ClassAntineoplastics; Small molecules; Vascular disorder therapies
Mechanism of ActionProto-oncogene protein c-akt inhibitors
Orphan Drug StatusYes - Hereditary haemorrhagic telangiectasia
Phase IVascular disorders
PreclinicalBreast cancer; Prostate cancer
No development reportedHereditary haemorrhagic telangiectasia
28 Dec 2025No recent reports of development identified for phase-I development in Hereditary haemorrhagic telangiectasia in Belgium (PO, Capsule)
28 Dec 2025No recent reports of development identified for phase-I development in Hereditary haemorrhagic telangiectasia in France (PO, Capsule)
28 Dec 2025No recent reports of development identified for phase-I development in Hereditary haemorrhagic telangiectasia in Italy (PO, Capsule)
In a 15 mL reaction tube was added 6-bromo-1-ethyl-7-phenyl-1H-pyrido[2,3-b][1,4]oxazin-2(3H)-one (50 mg, 0.150 mmol), tert-butyl ((1s,3s)-3-hydroxy-1-(4-(4,4,5,5-tetramethyl-1,3,2-dioxaborolan-2-yl)phenyl)cyclobutyl)carbamate (49 mg, 0.125 mmol) and cesium carbonate (204 mg, 0.625 mmol) in a mixture of 1,4-dioxane (2.3 ml) and water (0.8 ml) to give a colourless solution. This was degassed by bubbling nitrogen for 15 minutes, followed by the addition of [1,1'-bis(diphenylphosphino)ferrocene]dichloropalladium(II) dichloromethane adduct (20 mg, 0.025 mmol) and degassing for a further 5 minutes. The reaction mixture was heated to 50°C under a nitrogen atmosphere for one hour then allowed to cool to room temperature, diluted with water (5 ml) and extracted into ethyl acetate (3 x 5 ml). The combined organic phases were dried over Na2SO4, filtered and concentrated to dryness under reduced pressure. The residue was purified by Biotage chromatography (cyclohexane:ethyl acetate, gradient elution from 90:10 to 0:100) to give the desired product as an off-white solid (45 mg, 70% yleld). Ή-NMR (500 MHz, CDCl3) δ 7.29-7.35 (5H, m), 7.28 (1H, s), 7.18-7.24 (4H, m), 4.96 (1H, br s), 4.88 (2H, s), 4.05 (1H, br s), 4.01 (2H, q), 2.98 (2H, br s), 2.75 (2H, br s), 1.20-1.51 (9H, br m), 1.32 (3H, t). LCMS (Method D) RT = 1.25 min, M+H+ = 516.20.
tert-butyl ((1s,3s)-1-(4-(1-ethyl-2-oxo-7-phenyl-2,3-dihydro-1H-pyrido[2,3-b][1,4]oxazin-6-yl)phenyl)-3-hydroxycyclobutyl)carbamate (45 mg, 0.087 mmol) was dissolved in TFA (1 mL) and stirred for 30 seconds. The solution was immediately concentrated to dryness under reduced pressure. The residue was dissolved in diethyl ether (~3 mL) and concentrated to dryness under reduced pressure three times. The residue was then slurried in diethyl ether (3 mL) and after settling the supernatant solvent removed by pipette. This was repeated three times. The remaining solvent was removed by freeze drylng overnight to give the desired compound as an off-white solid (33 mg, 71% yleld).
1H-NMR (500 MHz, MeOD) δ 7.55 (1H, s), 7.39-7.42 (4H, m), 7.27-7.31 (3H, m), 7.20-7.24 (2H, m), 4.93 (2H, s), 4.01-4.11 (3H, m), 3.03-3.11 (2H, m), 2.42-2.50 (2H, m), 1.28 (3H, t). LCMS (Method D) RT = 0.74 min, M+H+ = 416.20.
Example 1: Synthesis of 6-(4-(l-amino-3-hvdroxycvclobutyl)phenyl)-l-ethyl-7-phenyl-lH-pyrido[2,3-b][l,4]oxazin-2(3H)-one
6-(4-(l-amino-3-hydroxycyclobutyl)phenyl)-l-ethyl-7-phenyl-lH-pyrido[2,3-b][l,4]oxazin-2(3H)-one (referred to herein as "VAD044 free base") was synthesized in accordance with the protocol as set out in W02011077098 - see in particular Examples 97, 113 and 139 (reproduced below):
Synthesis of 6-(4-((ls,3s)-l-amino-3-hvdroxycvclobutyl)phenyl)-l-ethyl-7-phenyl-lH-pyrido[2,3-bHl,41oxazin-2(3H)-one: from WO2Q11077098 Example 139:
In a 15 mL reaction tube was added 6-bromo-l-ethyl-7-phenyl-lH-pyrido[2,3-b][l,4]oxazin-2(3H)-one* (50 mg, 0.150 mmol), tert-butyl((ls,3s)-3-hydroxy-l-(4-(4,4,5,5-tetramethyl-l,3,2-dioxaborolan-2-yl)phenyl)cyclobutyl)carbamate** (49 mg, 0.125 mmol) and cesium carbonate (204 mg, 0.625 mmol) in a mixture of 1,4-dioxane (2.3 ml) and water (0.8 ml) to give a colourless solution. This was degassed by bubbling nitrogen for 15 minutes, followed by the addition of [1,1'-bis(diphenylphosphino)ferrocene]dichloropalladium(ll) dichloromethane adduct (20 mg, 0.025 mmol) and degassing for a further 5 minutes. The reaction mixture was heated to 50°C under a nitrogen atmosphere for one hour then allowed to cool to room temperature, diluted with water (5 ml) and extracted into ethyl acetate (3 x 5 ml). The combined organic phases were dried over Na2SO4, filtered and concentrated to dryness under reduced pressure. The residue was purified by Biotage chromatography (cyclohexane:ethyl acetate, gradient elution from 90:10 to 0:100) to give the desired product as an off- white solid (45 mg, 70% yield). 1H-NMR (500 MHz, CDCI3) 6 7.29-7.35 (5H, m), 7.28 (1H, s), 7.18-7.24 (4H, m), 4.96 (1H, br s), 4.88 (2H, s), 4.05 (1H, br s), 4.01 (2H, q), 2.98 (2H, br s), 2.75 (2H, br s), 1.20-1.51 (9H, br m), 1.32 (3H, t). LCMS (Method D) RT = 1.25 min, M+H+ = 516.20.
tert-butyl((ls,3s)-l-(4-(l-ethyl-2-oxo-7-phenyl-2,3-dihydro-lH-pyrido[2,3-b][l,4]oxazin-6-yl)phenyl)-3- hydroxycyclobutyl)carbamate (45 mg, 0.087 mmol) was dissolved in TFA (1 mL) and stirred for 30 seconds. The solution was immediately concentrated to dryness under reduced pressure. The residue was dissolved in diethyl ether (~3 mL) and concentrated to dryness under reduced pressure three times. The residue was then slurried in diethyl ether (3 mL) and after settling the supernatant solvent removed by pipette. This was repeated three times. The remaining solvent was removed by freeze drying overnight to give the desired compound as an off-white solid (33 mg, 71% yield). 1H-NMR (500 MHz, MeOD) 6 7.55 (1H, s), 7.39- 7.42 (4H, m), 7.27-7.31 (3H, m), 7.20- 7.24 (2H, m), 4.93 (2H, s), 4.01-4.11 (3H, m), 3.03-3.1 1 (2H, m), 2.42-2.50 (2H, m), 1.28 (3H, t). LCMS (Method D) RT = 0.74 min, M+H+ = 416.20.
To a suspension of sodium hydride (5.31 g, 133 mmol) in 1,4-dioxane (250 ml), ethyl glycolate (12.56 ml,
133 mmol) was added drop wise over a period of 30 minutes ensuring that the temperature was maintained below 30°C. The resulting thick suspension was stirred at room temperature for 15 minutes.
In a separate II round- bottomed flask was added 5-bromo-2-chloro-3-nitropyridine (21 g, 88 mmol) in
1,4-dioxane (150 ml) to give a brown solution. The suspension of sodium hydride and ethyl glycolate was added drop wise over a period of 30 minutes at 0°C. The resulting reaction mixture was heated to 80°C overnight.
The reaction mixture was concentrated under reduced pressure and the crude residue was purified by
Biotage silica chromatography (gradient 0% to 10% ethyl acetate in n-hexanes) to give the title compound
In a II round-bottomed flask was added ethyl 2-(5-bromo-3-nitropyridin-2-yloxy)acetate (18.33 g, 60.1 mmol), phenylboronic acid (10.99 g, 90 mmol), triphenylphosphine (4.73 g, 18.02 mmol) and cesium fluoride (45.6 g, 300 mmol) in 1,2-dimethoxyethane (300 ml) to give a yellow solution. The reaction mixture was degassed by bubbling nitrogen for 30 minutes. Pallad ium (II ) acetate (2.023 g, 9.01 mmol) was added and the mixture was heated to 75°C under a nitrogen atmosphere overnight. The reaction mixture was allowed to cool to room temperature and concentrated to dryness under reduced pressure to give a brown solid. This was re-dissolved in dichloromethane, filtered and concentrated to dryness under reduced pressure to give a brown solid The crude residue was purified via Biotage chromatography (gradient 5% to 60% ethyl acetate in n-hexanes) to give the title compound (6.9g, 38%). 1H NMR (500 MHz, CDCI3) 6 8.58 (1H, s), 8.56 (1H, s), 7.59-7.52 (2H, m), 7.48-7.46 (2H, m), 7.45-7.43 (1H, m), 5.13 (2H, s), 4.30-4.26 (2H, q), 1.33-1.30 (3H, t).
In a 500 ml round-bottomed flask was added ethyl 2-(3-nitro-5-phenylpyridin-2-yloxy)acetate (4.6 g, 15.22 mmol) in hydrochloric acid, 37% (40 ml) to give a yellow suspension. The mixture was cooled to 0-5°C followed by the portion wise addition of tin powder (9.94 g, 84 mmol). The addition proved to be exothermic. Caution should be taken while adding. The mixture was then stirred at room temperature for further 30 minutes until all foaming ceased. The reaction mixture was heated to 80°C under a nitrogen atmosphere for 3 hours. The reaction mixture cooled to room temperature and diluted with water (800ml). The white precipitate was isolated by filtration, washed with water (100 ml) and sucked dry to give a white solid. The solid was azeotroped with toluene (3 x 30 ml) to give a white solid as the title compound (2.6g, 77%). XH NMR (500 MHz, (CD3)2SO) 6 10.41 (1H, s), 8.10 (1H, s), 7.59 (2H, d), 7.49-7.42 (2H, t), 7.39-7-38 (1H, d), 4.83 (2H, s).
In a 10ml microwave vial was 7-phenyl-l H-pyrido[2,3-b][l ,4]oxazin-2(3H)-one (50 mg, 0.221 mmol) and N-bromosuccinimide (78.6 mg, 0.441 mmol) in dimethylformamide (1 ml). The reaction mixture was heated to 80°C under microwave irradiation for 30 minutes. The reaction mixture was cooled to room
temperature and diluted with ethyl acetate (10ml). The organic solution was washed with water (2x10ml) and brine (2x10ml). The organic phase was dried over anhydrous Na2SO4, filtered and concentrated under reduced pressure. The crude residue was purified via Biotage chromatography (gradient 0% to 5% methanol in dichloromethane) to give the title compound as a yellow solid (61 mg, 90%). 1H NMR (500 MHz, CD3OD) 6 7.48-7.32 (5H, m), 7.12 (1 H, s), 4.82 (2H, s).
In a 15 mL reaction tube was added 6-bromo-7-phenyl-lH-pyrido[2,3-b][l,4]oxazin-2(3H)-one (300 mg, 0.983 mmol), iodoethane (0.095 mL, 1.180 mmol) and potassium carbonate (408 mg, 2.95 mmol) in anhydrous N,N-dimethylformamide (1 mL) to give a brown suspension. This was stirred at 50 °C under a nitrogen atmosphere for 60 minutes. The reaction mixture was diluted with saturated sodium bicarbonate solution (5 mL) and extracted into ethyl acetate (3 x 5 mL). The combined organic phases were washed with 50:50 water.brine (3 x 5 mL), dried over Na2SO4, filtered and concentrated to dryness under reduced pressure to give a brown solid. This was purified by Biotage chromatography (25g silica cartridge, cyclohexane:ethyl acetate, gradient elution from 90:10 to 20:80) to give the title compound as a beige solid (160 mg, 48.8 % yield). XH NMR (500 MHz, CDCI3) 6 7.58-7.37 (5H, m), 7.21 (1H, s), 4.86 (2H, s), 3.96 (2H, q), 1.27 (3H, t). LCMS (Method D) RT 1.293 min, M+l= 334.
n a 40 mL reaction tube was added tert-butyl(ls,3s)-l-(4-bromophenyl)-3- hydroxycyclobutylcarbamate*** (0.25 g, 0.731 mmol) in anhydrous tetrahydrofuran (14 ml) to give a colourless solution. This was degassed by bubbling nitrogen for 20 minutes, followed by the addition of [l,l'-bis(diphenylphosphino)ferrocene]dichloropalladium(ll) dichloromethane adduct (60 mg, 0.073 mmol). After bubbling nitrogen for a further 15 minutes, potassium acetate (143 mg, 1.461 mmol) and bis(pinacolato)diboron (223 mg, 0.877 mmol) were added. The reaction mixture was heated to reflux overnight then concentrated to dryness under reduced pressure and purified by Biotage chromatography (cyclohexane:ethyl acetate, gradient elution from 88:12 to 0:100) to give the desired product as a colourless oil that solidified upon standing (240 mg, 84% yield). 1H-NMR (500 MHz, CDCI3) 6 7.71 (2H, d), 7.44 (2H, d), 4.15 (1H, br s), 2.87-2.98 (2H, m), 2.27-2.44 (2H, m), 1.22-1.49 (21H, br m).
(*** synthesis described in WO2009148887 and WO2009148916)
Enozertinib (formerly ORIC-114) is an investigational, orally bioavailable, and brain-penetrant dual EGFR/HER2 inhibitor developed by ORIC Pharmaceuticals. It targets cancers with exon 20 insertion and atypical EGFR mutations. Its core profile highlights its ability to cross the blood-brain barrier.
How it Works
Enozertinib acts as an irreversible, mutant-selective covalent inhibitor. By blocking overactive EGFR and HER2 signaling, it induces cell death and inhibits tumor growth. Because it penetrates the central nervous system (CNS), it is uniquely suited to treat both primary brain tumors and brain metastases—a common complication in non-small cell lung cancer (NSCLC).
Enozertinib is an orally bioavailable, central nervous system (CNS) penetrating, mutant-selective covalent inhibitor of epidermal growth factor receptor (EGFR; ErbB1) and human epidermal growth factor receptor 2 (HER2; EGFR2; ErbB2) alterations, including exon 20 insertion (Ex20ins) mutations, with potential antineoplastic activity. Upon oral administration, enozertinib selectively targets, irreversibly binds to and inhibits the activity of EGFR or HER2 insertions or mutations. This prevents EGFR/HER2-mediated signaling. This may induce cell death and inhibit tumor growth in EGFR/HER2-overexpressing tumor cells. Enozertinib is able to penetrate the blood-brain-barrier (BBB) and may therefore exert its activity against EGFR Ex20ins-driven CNS primary tumors and CNS metastases. The ErbB receptor tyrosine kinase family is involved in key cellular functions, including cell growth and survival. EGFR and HER2 alterations constitutively upregulate kinase activity.
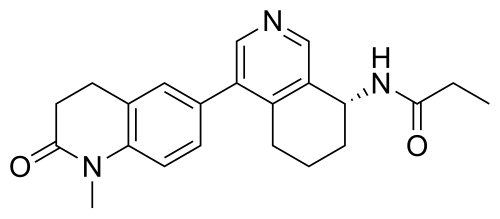
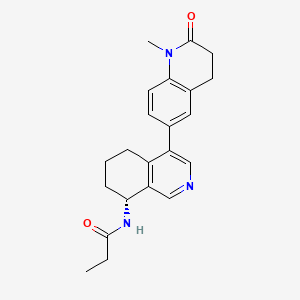
Baxdrostat is under investigation in clinical trial NCT06344104 (A Phase III Study to Investigate the Efficacy and Safety of Baxdrostat in Asian Participants With Uncontrolled Hypertension on Two or More Medications Including Participants With Resistant Hypertension).
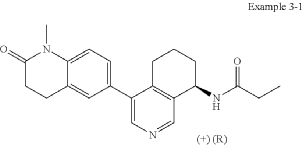
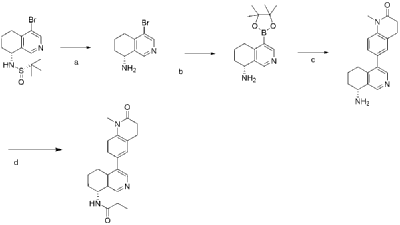
In analogy to the procedures described for the preparation of intermediate A-2 [E] and for the preparation of intermediate B-1, Suzuki reaction of (+)-(R)-4-bromo-5,6,7,8-tetrahydroisoquinolin-8-amine (intermediate B-3b) with 1-methyl-6-(4,4,5,5-tetramethyl-[1,3,2]dioxaborolan-2-yl)-3,4-dihydro-1H-quinolin-2-one (intermediate A-1) gave (R)-6-(8-amino-5,6,7,8-tetrahydroisoquinolin-4-yl)-1-methyl-3,4-dihydroquinolin-2(1H)-one and after subsequent reaction with propionyl chloride the title compound as colorless solid. MS: 364.2 (M+H +).
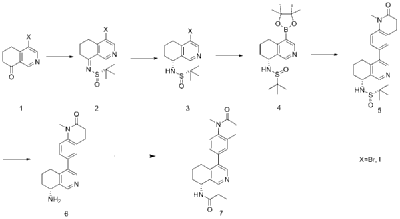
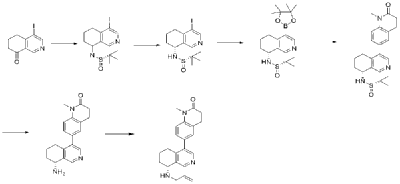
Dissolve 4-bromo-6,7-dihydroisoquinolin-8(5H)-one (1.56 g, 6.9 mmol) and (S)-tert-butylsulfenamide (2.51 g, 20.7 mmol) in 20 mL of tetrahydrofuran. Add ethyl titanate (10.08 mL, 48.28 mmol). Heat to 65°C and stir for 48 hours. Cool to room temperature, add ethyl acetate and water, stir for 15 minutes, and remove the resulting solid by filtration. Separate the liquids, dry the organic phase over anhydrous sodium sulfate, filter, and evaporate to dryness under reduced pressure to obtain the crude product (S,Z)-N-(4-bromo-6,7-dihydroisoquinolin-8(5H)-tert-butylsulfenimide), which is used directly in the next step.
Step B
Compound (S,Z)-N-(4-bromo-6,7-dihydroisoquinoline-8(5H)-tert-butylsulfonyl imide) (1.98 g, 6 mmol) was dissolved in 15 mL of tetrahydrofuran and cooled to -45°C. Sodium borohydride (0.34 g, 9.0 mmol) was added, and the mixture was allowed to return to room temperature and stirred for 18 hours. The mixture was quenched with ice water and extracted with dichloromethane. The resulting organic phase was washed with saturated brine, dried over anhydrous sodium sulfate, and evaporated to dryness under reduced pressure. The residue was purified by column chromatography to obtain compound (S)-N-(4-bromo-6,7-dihydroisoquinoline-8(5H))-tert-butylsulfonyl imide (755 mg, 38% yield). LC/MS (ESI): m/z = 331.2 [M+H] + .
Step C
To a mixture of (S)-N-(4-bromo-6,7-dihydroisoquinoline-8(5H))-tert-butylsulfonimide (0.66 g, 2 mmol), pinacol diboronate (1.05 g, 2.1 mmol), and AcOK (0.578 g, 6 mmol) in toluene (10 mL) was added Pd(dppf)Cl 2 (0.144 g, 0.2 mmol). The mixture was degassed and stirred at 130 ° C for 3 hours. The reaction mixture was filtered and concentrated to give a residue. EtOAc (15 mL) and water (10 mL) were added to the residue. The organic phase was washed with brine (50 mL), dried over anhydrous sodium sulfate, filtered and concentrated to give a residue. The residue was purified by column chromatography (SiO 2 ) and eluted with 30-40% ethyl acetate in petroleum ether to afford (S)-N-tert-butylsulfonamido-6,7-dihydroisoquinolin-8(5H)-4-boronic acid pinacol ester (0.45 g, 60% yield). LC/MS (ESI): m/z = 378.3 [M+H] + .
Step D
To a reaction flask, add 6-bromo-1-methyl-3,4-dihydroquinolin-2(1H)-one (0.29 g, 1.2 mmol), (S)-N-tert-butylsulfonamido-6,7-dihydroisoquinolin-8(5H)-4-boronic acid pinacol ester (0.42 g, 1.26 mmol), bistriphenylphosphine palladium dichloride (84 mg, 0.12 mmol), cuprous iodide (38 mg, 0.2 mmol), triethylamine (1.01 g, 10.0 mmol), and 15 mL of N,N-dimethylformamide. The atmosphere was purged with nitrogen three times and the reaction was stirred at 90°C overnight. After cooling to room temperature, the reaction mixture was diluted with ethyl acetate and water, and extracted with ethyl acetate. The resulting organic phase was washed with water and saturated brine, dried over anhydrous sodium sulfate, and evaporated to dryness under reduced pressure. The residue was purified by column chromatography to afford (S)-2-methyl-N-((R)-4-(1-methyl-2-oxo-1,2,3,4-tetrahydroquinolin-6-yl)-5,6,7,8-tetrahydroisoquinolin-8-yl)tert-butylsulfonimide (0.37 g, 74% yield) as a yellow solid. LC/MS (ESI): m/z = 411.5 [M+H] + .
Step E
Compound (S)-2-methyl-N-((R)-4-(1-methyl-2-oxo-1,2,3,4-tetrahydroquinolin-6-yl)-5,6,7,8-tetrahydroisoquinolin-8-yl)tert-butylsulfonimide (0.33 g, 0.80 mmol) was dissolved in 1 mL of dichloromethane, and 1 mL of trifluoroacetic acid was added. The mixture was stirred and reacted for 1 hour. The reaction solution was concentrated under reduced pressure. The residue was purified by reverse preparative column chromatography to obtain compound (R)-6-(8-amino-5,6,7,8-tetrahydroisoquinolin-4-yl)-1-methyl-3,4-dihydroquinolin-2(1H)-one (0.24 g, 97% yield). LC/MS (ESI): m/z = 307.1 [M+H] + .
Step F
To a reaction flask, add (R)-6-(8-amino-5,6,7,8-tetrahydroisoquinolin-4-yl)-1-methyl-3,4-dihydroquinolin-2(1H)-one (100 mg, 0.33 mmol), triethylamine (51 mg, 0.5 mmol), and 4 ml of tetrahydrofuran. After cooling in an ice-water bath, slowly add a solution of propionyl chloride (46.25 mg, 0.5 mmol) in 0.5 ml of tetrahydrofuran dropwise. Stirring is continued for 4 hours after addition. The reaction mixture is quenched with methanol and evaporated to dryness under reduced pressure. The residue is purified by column chromatography to obtain the target compound, Baxdrostat (46 mg, 38% yield). LC/MS(ESI):m/z=363.1[M+H]+.H NMR(400MHz, CDCl3)ppm 1.22(t,3H)1.79(s,3H)2.07(s,1H)2.28(q,2H)2.43-2.68(m,2H)2.71(t,2H)2.82-3.12(m,2H) 3.40(s,3H)5.34(d,1H)5.78(d,1H)7.05(d,1H)7.09(s,1H)7.17(d,1H)8.28(s,1H)8.49(s,1H)
Example 2
Step A
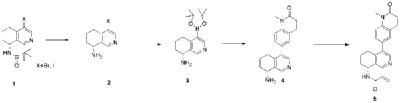
Compound (S)-N-(4-bromo-6,7-dihydroisoquinolin-8(5H))-tert-butylsulfonylimide (1.65 g, 5 mmol) was dissolved in 20 mL of dichloromethane, and 20 mL of trifluoroacetic acid was added. The mixture was stirred and reacted for 1 hour. The reaction solution was concentrated under reduced pressure. The residue was purified by reverse-phase preparative column chromatography to obtain compound (R)-4-bromo-5,6,7,8-tetrahydroisoquinolin-8-amine (1.07 g, 94% yield). LC/MS (ESI): m/z = 226.0 [M+H] + .
Step B
To a mixture of (R)-4-bromo-5,6,7,8-tetrahydroisoquinolin-8-amine (0.86 g, 3.8 mmol), pinacol diboron (2 g, 4 mmol), AcOK (1.10 g, 11.4 mmol) in toluene (10 mL) was added Pd(dppf)Cl 2 (0.27 g, 0.38 mmol). The mixture was degassed and stirred at 130 ° C for 3 hours. The reaction mixture was filtered and concentrated to give a residue. EtOAc (10 mL) and water (10 mL) were added to the residue. The organic phase was washed with brine (10 mL), dried over anhydrous sodium sulfate, filtered and concentrated to give a residue. The residue was purified by column chromatography (SiO 2 ) and eluted with 30-40% ethyl acetate in petroleum ether to afford (R)-8-amino-5,6,7,8-tetrahydroisoquinoline-4-boronic acid pinacol ester (0.68 g, 65% yield). LC/MS (ESI): m/z = 274.1 [M+H] + .
Step C
To a reaction flask, add 6-bromo-1-methyl-3,4-dihydroquinolin-2(1H)-one (0.72 g, 3.0 mmol), (R)-8-amino-5,6,7,8-tetrahydroisoquinolin-4-boronic acid pinacol ester (0.99 g, 3.6 mmol), bistriphenylphosphine palladium dichloride (210 mg, 0.3 mmol), and potassium phosphate monohydrate (204 mg, 0.9 mmol). Dissolve the mixture in dioxane and water (9:1, 30 mL). Replace the atmosphere with nitrogen three times and allow the mixture to react overnight at 90°C with stirring. Cool to room temperature, dilute the reaction solution with ethyl acetate and water, and extract with ethyl acetate. The resulting organic phase is then washed with water and saturated brine, dried over anhydrous sodium sulfate, and evaporated to dryness under reduced pressure. The residue was purified by column chromatography to obtain (R)-6-(8-amino-5,6,7,8-tetrahydroisoquinolin-4-yl)-1-methyl-3,4-dihydroquinolin-2(1H)-one (0.81 g, 88% yield). LC/MS (ESI): m/z = 307.1 [M+H] + . The target compound, Baxdrostat, was then prepared using a method similar to the last step in Example 1.
Example 3
Step A
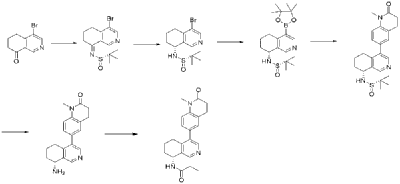
4-Bromo-6,7-dihydroisoquinolin-8(5H)-one (1.88 g, 6.9 mmol) and (S)-tert-butylsulfenamide (2.51 g, 20.7 mmol) were dissolved in 20 mL of tetrahydrofuran. Ethyl titanate (10.08 mL, 48.28 mmol) was added and the mixture was heated to 65°C with stirring for 48 hours. After cooling to room temperature, ethyl acetate and water were added and stirred for 15 minutes. The resulting solid was removed by filtration. The organic phase was separated and dried over anhydrous sodium sulfate, filtered, and evaporated to dryness under reduced pressure to obtain the crude product (S,Z)-N-(4-bromo-6,7-dihydroisoquinolin-8(5H)-tert-butylsulfenimide), which was used directly in the next step. LC/MS (ESI): m/z = 376.2 [M+H] + .
Step B
Compound (S,Z)-N-(4-iodo-6,7-dihydroisoquinoline-8(5H)-tert-butylsulfonyl imide) (2.26 g, 6 mmol) was dissolved in 15 mL of tetrahydrofuran and cooled to -45°C. Sodium borohydride (0.36 g, 9.0 mmol) was added, and the mixture was allowed to return to room temperature and stirred for 18 hours. The mixture was quenched with ice water and extracted with dichloromethane. The resulting organic phase was washed with saturated brine, dried over anhydrous sodium sulfate, and evaporated to dryness under reduced pressure. The residue was purified by column chromatography to obtain compound (S)-N-(4-iodo-6,7-dihydroisoquinoline-8(5H))-tert-butylsulfonyl imide (1.04 g, 46% yield). LC/MS (ESI): m/z = 378.0 [M+H] + .
Step C
To a mixture of (S)-N-(4-iodo-6,7-dihydroisoquinoline-8(5H))-tert-butylsulfonimide (0.76 g, 2 mmol), pinacol diboronate (1.05 g, 2.1 mmol), and AcOK (0.578 g, 6 mmol) in toluene (10 mL) was added Pd(dppf)Cl 2 (0.144 g, 0.2 mmol). The mixture was degassed and stirred at 130 ° C for 3 hours. The reaction mixture was filtered and concentrated to give a residue. EtOAc (15 mL) and water (10 mL) were added to the residue. The organic phase was washed with brine (50 mL), dried over anhydrous sodium sulfate, filtered and concentrated to give a residue. The residue was purified by column chromatography (SiO 2 ) and eluted with 30-40% ethyl acetate in petroleum ether to afford (S)-N-tert-butylsulfonamido-6,7-dihydroisoquinolin-8(5H)-4-boronic acid pinacol ester (0.51 g, 68% yield). LC/MS (ESI): m/z = 378.2 [M+H] + .
The next three steps were carried out in the same manner as in Example 1 to prepare the target compound Baxdrostat.
A new treatment has been shown to significantly lower blood pressure in people whose levels stay dangerously high, despite taking several existing medicines, according to the results of a Phase III clinical trial led by a UCL Professor. Globally, around 1.3 billion people have high blood pressure (hypertension), and in around half of cases the condition is uncontrolled or treatment resistant. These individuals face a much greater risk of heart attack, stroke, kidney disease, and early death. In the UK the number of people with hypertension is around 14 million.
The international BaxHTN trial, led by Professor Bryan Williams (UCL Institute of Cardiovascular Science), assessed the new drug baxdrostat—which is taken as a tablet—with participation from nearly 800 patients across 214 clinics worldwide.
The trial results showed that, after 12 weeks, patients taking baxdrostat (1 mg or 2 mg once daily in pill form) saw their blood pressure fall by around 9-10 mmHg more than placebo—a reduction large enough to cut cardiovascular risk. About four in 10 patients reached healthy blood pressure levels, compared with fewer than two in 10 on placebo.
Principal Investigator, Professor Williams, who is presenting the results at ESC, said, "Achieving a nearly 10 mmHg reduction in systolic blood pressure with baxdrostat in the BaxHTN Phase III trial is exciting, as this level of reduction is linked to substantially lower risk of heart attack, stroke, heart failure and kidney disease."
How baxdrostat works
Blood pressure is strongly influenced by a hormone called aldosterone, which helps the kidneys regulate salt and water balance.
Some people produce too much aldosterone, causing the body to hold onto salt and water. This aldosterone dysregulation pushes blood pressure up and makes it very difficult to control.
Addressing aldosterone dysregulation has been a key effort in research over many decades, but it has been so far difficult to achieve.
Baxdrostat works by blocking aldosterone production, directly addressing this driver of high blood pressure (hypertension).
Professor Williams, Chair of Medicine at UCL, said, "These findings are an important advance in treatment and in our understanding of the cause of difficult-to-control blood pressure.
"Around half of people treated for hypertension do not have it controlled, however this is a conservative estimate and the number is likely higher, especially as the target blood pressure we try to reach is now much lower than it was previously.
"In patients with uncontrolled or resistant hypertension, the addition of baxdrostat 1mg or 2mg once daily to background antihypertensive therapy led to clinically meaningful reductions in systolic blood pressure, which persisted for up to 32 weeks with no unanticipated safety findings.
"This suggests that aldosterone is playing an important role in causing difficult to control blood pressure in millions of patients and offers hope for more effective treatment in the future."
Historically, higher-income Western countries were reported to have far higher levels of hypertension. However, largely due to changing diets (adding less salt to food), the numbers of people living with the condition is now far higher in Eastern and lower-income countries. More than half of those affected live in Asia, including 226 million people in China and 199 million in India.
Professor Williams added, "The results suggest that this drug could potentially help up to half a billion people globally—and as many as 10 million people in the UK alone, especially at the new target level for optimal blood pressure control."
AS ON JUNE2025 4.45 LAKHS VIEWS ON BLOG WORLDREACH AVAILABLEFOR YOUR ADVERTISEMENT
Dogra S, Shah S, Gitzel L, Pusukur B, Sood A, Vyas AV, Gupta R (July 2023). "Baxdrostat: A Novel Aldosterone Synthase Inhibitor for Treatment Resistant Hypertension". Current Problems in Cardiology. 48 (11): 101918. doi:10.1016/j.cpcardiol.2023.101918. PMID37399857. S2CID259320969.
Awosika A, Cho Y, Bose U, Omole AE, Adabanya U (October 2023). "Evaluating phase II results of Baxdrostat, an aldosterone synthase inhibitor for hypertension". Expert Opinion on Investigational Drugs. 32 (11): 985–995. doi:10.1080/13543784.2023.2276755. PMID37883217. S2CID264517675.